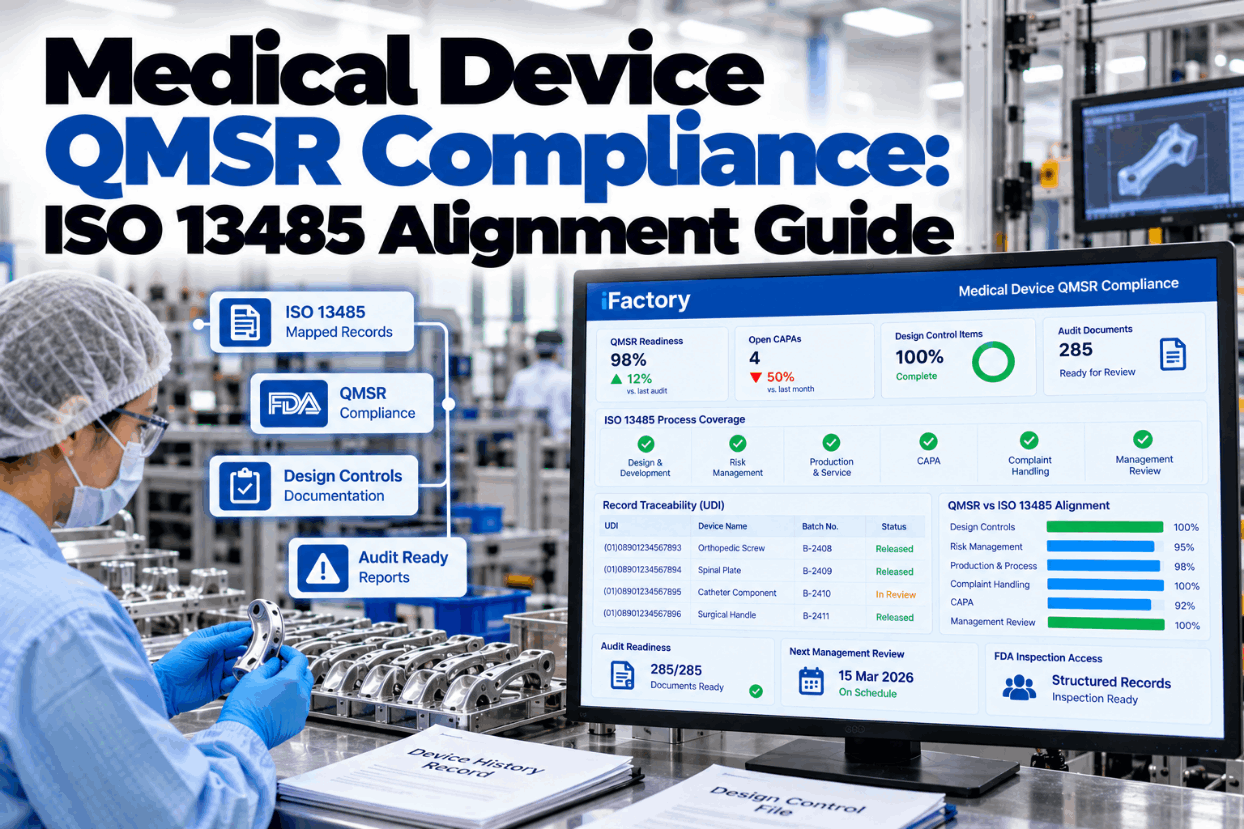
On February 2, 2026, FDA's Quality System regulation gave way to the Quality Management System Regulation. 21 CFR 820 now incorporates ISO 13485:2016 by reference, and records that investigators could not ask for before, such as management reviews and internal audits, are now open to inspection. This guide explains what changed, what stayed, and how to keep the records ready. To see how a records toolkit maps to ISO 13485, book a short QMSR records demo.
Keep Every QMSR Record One Click From the Investigator
iFactory QMSR Toolkit maps your quality records to ISO 13485:2016 clauses, links complaints to UDI and MDR decisions, and keeps management review and audit reports organized for FDA inspection access.
What Happened on February 2, 2026
FDA published the final rule on February 2, 2024 and gave manufacturers two years to prepare. On the effective date, the rule renamed the regulation and replaced most of the old Part 820 text with a requirement to follow ISO 13485:2016, plus Clause 3 of ISO 9000:2015 for vocabulary. FDA also stopped using the Quality System Inspection Technique (QSIT) that day and moved to Compliance Program 7382.850. FDA's FAQ notes that investigators may still review records made before that date. Check your own position with our quality team.
A Crosswalk From the Old QS Regulation to ISO 13485:2016
Most of the work is mapping. Requirements you already meet under the old sections usually have a home in an ISO 13485 clause, but the wording, the record names and sometimes the depth differ. The crosswalk below is a starting point for a gap check, not a legal opinion, so confirm each line against the standard and your own procedures. Walk through your mapping in a crosswalk session.
Old Words, New Homes
FDA retired several familiar record names because ISO 13485 organizes the same content differently. The device master record, design history file and device history record are no longer QMSR terms. The content still has to exist, but it now sits in the files ISO 13485 describes, and definitions come mainly from ISO 9000:2015 Clause 3, with FDA exceptions in 820.3. Ask our support team how to relabel records without rebuilding them.
Three Record Sets That Are No Longer Off Limits
Under the old rule, section 820.180(c) kept management reviews, quality audit reports and supplier audit reports out of routine FDA inspection. FDA states plainly that these exceptions are not kept in the QMSR, and that the records should be readily available on inspection. That changes how they should be written: clear, factual and complete, with actions tracked to closure. See how review packs are organized in a records walkthrough.
Section 820.35(d) lets you mark records you consider confidential. That helps FDA decide what may be disclosed under 21 CFR part 20. It does not keep the record from an investigator.
Answer the Records Request Before the Coffee Gets Cold
iFactory QMSR Toolkit keeps each record tagged to its ISO 13485 clause, links complaints to UDI and MDR decisions and shows what is ready, what is missing and who owns it.
ISO 13485 Is the Base, Not the Whole Rule
The QMSR is ISO 13485:2016 plus a set of FDA requirements on top. Section 820.10 ties the standard to other parts of FDA's rules, including medical device reporting, corrections and removals, tracking and UDI. Section 820.35 adds record content, and 820.45 keeps labeling and packaging controls. A plant that is certified to ISO 13485 alone can still miss these. Review your FDA-specific layer with our regulatory support team.
How FDA Now Organizes a Device Inspection
Compliance Program 7382.850 replaces the four QSIT subsystems with six QMS areas and four other applicable FDA requirements. Investigators look at elements within each area, and risk to patients and users guides where they dig deeper. In practice that means complaint trends, reportable events and recalls can steer an inspection toward the processes behind them. Management oversight now includes the reviews and audits that were once exempt. Prepare your area owners in a readiness session.
A Complaint Record That Meets 820.35(a)
Complaint records are where ISO 13485 and FDA's own rules meet. Section 820.35(a) lists seven items every complaint record must hold, and complaints that must be reported under Part 803 need extra detail on top. Use the fields below as a template, and add your own investigation and trend fields. Request an editable version from our support team.
The Shop-Floor Records That Carry the Most Weight
Quality owns the system, but much of the evidence is made on the floor. Production and Service Provision is one of the six inspection areas, and the records behind it come from maintenance, metrology and operations. Plant managers who keep these complete make every other part of an inspection easier. Review your floor records with our plant quality specialists.
Five Steps From Gap Check to Inspection-Ready
If the transition work is not finished, or was done in a hurry before the deadline, these steps are a practical order. Each one produces something an investigator can see, which keeps the effort focused on evidence rather than paperwork for its own sake. FDA's FAQ suggests a comparative analysis showing that records made before the effective date also meet the QMSR, so build that into step two. Plan your steps on a support call.
Sizing the Time Spent Finding Records
The clearest cost to size is the time it takes to pull records during an inspection or audit. Every input below is an assumption to show the arithmetic, so replace each with your own figures. Our team can help you build the full case.
QMSR Toolkit and AI Server in One Package
iFactory delivers the QMSR Toolkit as a turnkey bundle. A pre-configured NVIDIA AI server ships racked and ready with the ISO 13485 clause map, record connectors and search pre-loaded, so quality records stay on site under your control. You rack it, plug in power and Ethernet, and the AI is live. To scope it for your plant, request a turnkey quote.
- Pre-configured NVIDIA AI edge server, racked and ready
- ISO 13485:2016 clause map for your records
- Complaint, UDI and MDR decision links
- Management review and audit report library
- Connectors to your QMS, ERP and CMMS records
- Inspection view with readiness and owners
- 24×7 remote monitoring of software and hardware
Server installed, record sources linked, procedures and records mapped to clauses.
Missing records flagged to owners, complaint and UDI links checked.
Inspection view switched on, teams trained, mock inspection run.
A Records Layer Built for QMSR Inspections
iFactory sits across the systems where quality evidence already lives and organizes it the way the QMSR and FDA investigators expect. Book a pilot planning call to see it on your records.
What Quality and Plant Managers Ask About the QMSR
Every Record Mapped, Every Request Answered
Map records to ISO 13485, link complaints to UDI and MDR decisions and keep reviews and audits ready for FDA. The full system is delivered turnkey and goes live in 6 to 12 weeks.